This guide explains Pmoc Abrava in practical, industry-focused terms—what it is, where it appears in specialized chemistry workflows, and how professionals manage quality, traceability, and safety during handling. Background information is provided objectively on the keyword context, typical regulatory expectations, and decision factors used by procurement and technical teams when evaluating this input.
In specialized chemical development and manufacturing, Pmoc Abrava is referenced as a defined chemical building block or intermediate used to support downstream synthesis steps. In many professional environments—ranging from discovery labs to pilot plant and scale-up groups—the term functions less like a “generic reagent” and more like a process-sensitive input whose quality characteristics can materially affect reaction outcomes. From an industry perspective, the very important point is not just “what it is,” but how it behaves when processed inside controlled workflows: purity requirements, documentation quality, compatibility with solvents and reagents, and the reliability of supply. When technical teams evaluate Pmoc Abrava, they generally focus on reproducibility—ensuring that every batch performs predictably in reaction planning, purification, and final conversion steps.
Because procurement outcomes depend on more than price, professionals typically validate multiple layers of assurance. Instead of relying on a catalog line alone, they seek evidence that the material matches the real needs of the specific step. The most common categories include: certificate-of-analysis (CoA) alignment, analytical method transparency, storage stability, and track-and-trace records. Even when two suppliers quote similar specifications on paper, the practical difference often lies in documentation rigor, consistency between lots, and the ability to investigate deviations if a batch behaves unexpectedly.
It’s also typical that cross-functional stakeholders weigh in: chemists focus on performance attributes and impurity effects; quality assurance focuses on how those attributes are verified and documented; and procurement focuses on lead times and supply continuity. In regulated contexts, regulatory/compliance teams add additional expectations around audit trails, change control, and qualification status. Therefore, Pmoc Abrava in professional workflows often sits at the intersection of chemical understanding and quality-system discipline.
Note: Your request included placeholders such as price information and supplier details, but no concrete values were provided. Accordingly, this article avoids inventing pricing or naming unverified suppliers. Instead, it outlines the exact categories of information you should confirm with any vendor offering Pmoc Abrava so you can compare offers objectively.
Teams handling Pmoc Abrava usually treat it as a process-sensitive input rather than a purely transactional purchase. In workflows involving protective-group chemistry, protected precursors, intermediate transformations, or any sequence where selectivity depends on fine chemical differences, small variations in impurity profile can influence yield, selectivity, and downstream purification burden. Consequently, qualification often starts with a technical question that is more precise than “does it meet the spec?” A more practical framing is: “Will this material behave the same way in our reaction system, on our equipment, under our conditions, within our acceptable risk boundaries?”
Therefore, qualification typically proceeds along a structured pathway. Many teams implement a risk-based approach rather than using the same depth of testing for every material. For Pmoc Abrava, risk may increase when the intermediate is used early (where impurities can propagate), when the step is highly sensitive to moisture or trace contaminants, or when downstream purification is expensive or capacity-limited. In contrast, risk may be lower if the step tolerates variability and the downstream process can robustly remove impurities.
In practice, engineers and quality managers want a supplier that supports a risk-based qualification approach, including incoming inspection procedures and stability considerations. That means not only providing a CoA but also enabling you to understand the basis of that CoA—what was tested, what method was used, what acceptance criteria were applied, and how consistent those criteria are across time.
The term Pmoc Abrava may appear in catalogs, technical communications, or internal documentation tied to complex synthesis routes. While the exact identity and use-case depends on the specific naming convention used by the documentation owner, the general industry pattern for molecules with protective-group or intermediate roles includes:
From a compliance viewpoint, suppliers and users generally operate within expectations aligned to good documentation practices and, where applicable, GMP/GLP-style requirements. When chemical inputs are used in regulated contexts—such as pharmaceutical development, clinical manufacturing, or other quality-regulated industries—quality systems, change control, and batch traceability become central. In that environment, Pmoc Abrava is not just a chemical; it is a documented, traceable entity that must be linked to quality evidence over time.
When procuring Pmoc Abrava, the very valuable information is rarely the marketing description—it’s the verifiable technical documentation. Professionals typically request or review:
If a vendor provides strong documentation, labs often reduce friction during incoming quality checks. If documentation is vague, teams may spend time on additional testing—identity verification, impurity screening, or performance suitability—potentially delaying timelines and increasing operational costs. In a competitive development environment, documentation quality often becomes a hidden differentiator between suppliers.
Another professional point is that “specification sheet” and “CoA” can be different. The specification sheet is the contractual boundary; the CoA is the demonstrated result for a given lot. Teams usually focus on the CoA, but they also verify that the CoA uses a testing logic consistent with the spec (for example, the same impurity definitions, the same calculation basis, or comparable detection thresholds).
Even when a chemical is routinely used in a lab, handling still matters. For Pmoc Abrava, the practical handling principles used by experienced teams generally include:
Because you did not provide specific storage temperature or formulation details, this article stays general and professional. For any exact instruction set, always follow the SDS and supplier documentation for Pmoc Abrava. A professional handling approach also includes ensuring that staff training and facility controls match those recommendations—especially for moisture sensitivity and hazard controls.
In more mature production environments, teams also define how material is transferred (for example, whether it is transferred as supplied versus re-dispensed into secondary containers) and how secondary container labeling is managed. Those details seem administrative, but they strongly influence traceability and prevent mix-ups. For Pmoc Abrava, where small variability can matter, these operational details can be the difference between consistent performance and unexplained batch-to-batch drift.
Even if price quotes are available for Pmoc Abrava, seasoned procurement teams treat “cost” as a broader total delivered risk. The decision framework usually includes:
When internal stakeholders compare vendor proposals, they often ask: “How expensive is the downtime or extra testing if a batch fails our incoming criteria?” This reframes evaluation from purchasing to engineering economics. For instance, a supplier with cheaper unit pricing but a history of documentation gaps could shift costs into internal testing, delayed production runs, or—even worse—downstream yield loss and purification inefficiency.
Professional procurement also considers continuity. For multi-step synthesis campaigns, you may need sustained supply over multiple months. Teams typically ask for information about manufacturing capacity, batch scheduling stability, and whether the supplier can guarantee consistent specification and analytical methods across the planned time horizon.
The following table is designed to help you compare vendor documentation and qualification readiness for Pmoc Abrava. It does not include any links and it avoids assuming pricing or supplier identity. Use it as a checklist to ensure vendor comparisons are based on evidence rather than marketing language.
| Evaluation item | What a qualified supplier typically provides | Why it matters for Pmoc Abrava workflows | Conditions/requirements to confirm |
|---|---|---|---|
| Lot-specific CoA | Results tied to your lot/batch number, covering purity/impurity and any relevant physical or chemical tests | Supports incoming acceptance and reduces uncertainty about step-to-step reproducibility | Request CoA for the exact lot you will receive; verify test list matches your specification |
| Analytical method transparency | Method references or enough detail to interpret results (e.g., what impurity determinations are based on) | Helps your QA team understand how the numbers should be interpreted and repeated | Confirm whether methods are comparable to your internal requirements; plan verification testing as needed |
| Stability and storage guidance | Documented storage recommendations and shelf-life expectations under defined conditions | Degradation or moisture effects can influence impurity profile and performance | Confirm temperature guidance and recommended precautions; ensure your receiving/storage practices align |
| Traceability and labeling | Clear lot identifiers, batch records availability as appropriate, consistent packaging information | Supports audit readiness and troubleshooting if a batch underperforms | Ensure labels are readable and match the documentation; maintain receiving records |
| Quality agreement capability | Ability to support defined acceptance criteria and documented change control | Reduces risk of silent changes that affect reaction outcomes | Ask about change notification timelines and what documentation is included when changes occur |
| Safety documentation | Up-to-date SDS and compliant handling guidance | Supports correct lab controls and emergency planning | Review SDS before use; ensure your facility controls match recommended precautions |
To make this comparison practical, many teams also add internal columns (not shown in the table) such as “verified internally” and “risk rating.” In more mature programs, a supplier scorecard can also track responsiveness to deviation investigations, ability to provide historical CoA trends, and speed of issue resolution.
Below is a practical, step-by-step approach used by many technical teams to reduce risk when adopting Pmoc Abrava into an established workflow. Adapt the details to your internal policies and regulatory environment. The objective is to move from “we have a CoA” to “we can trust this material to perform consistently in our process, and we can explain why.”
Before ordering, identify which characteristics matter very for the step where Pmoc Abrava will be used. Common examples include impurity limits, physical form, and compatibility with your solvent system. If your process is sensitive, plan verification tests. Critical parameters are often those that influence:
Defining critical parameters also helps you decide how deep your qualification effort should be. If only a narrow set of attributes matters, you can limit testing to confirm those attributes rather than repeating exhaustive analysis.
Ask for a lot-specific CoA and the relevant safety documentation. Ensure the documentation corresponds to the same batch you will receive, not just a generic catalog specification. Many teams also request:
This step sets the baseline for incoming inspection and helps prevent misalignment between vendor reporting and your internal acceptance criteria.
At delivery, verify labeling, container integrity, and that the lot number matches the CoA. Then perform any internal checks your quality system requires (for instance, identity confirmation or basic purity screening). A receiving inspection typically includes:
Even if you plan to run a suitability test, receiving checks provide early detection of obvious issues and reduce the chance that you invest time in a material that is obviously out-of-spec.
Run a limited qualification reaction or process test at the scale needed to validate performance in your conditions. Track outcomes such as conversion, impurity formation patterns, and any purification shifts. This step is often the most persuasive evidence for “process suitability” because it demonstrates performance in your specific system.
When performing suitability tests, professional teams often define what success looks like beyond a simple yield number. For example, you may evaluate:
Because the intermediate might influence downstream steps, teams may also include minimal downstream checks (for example, a quick analytical readout after workup) to detect issues before scaling up.
If performance is consistent, formalize acceptance criteria for future batches. If performance varies, adjust supplier qualification scope or request additional evidence. Documenting results typically includes:
Acceptance criteria should be explicit and anchored to both CoA evidence and observed process performance. In some programs, acceptance criteria are expanded to include not just the vendor’s impurity thresholds but also internal process performance metrics.
Maintain batch records linking every use to lot numbers. Ask the supplier how they communicate changes in manufacturing processes that could impact Pmoc Abrava quality. Change control is important because even when a vendor remains within specification, underlying manufacturing changes can shift impurity spectra or alter physical properties.
Professional change control monitoring often includes:
If you do not have these mechanisms, you may discover changes only after you observe process drift—too late for a smooth transition.
Store according to documented guidance, and define internal timelines for “use before” conditions. For sensitive intermediates, disciplined handling is often as important as the initial specification. Stability discipline includes:
When these operational disciplines are enforced, Pmoc Abrava performance is more likely to match what you qualified during onboarding.
To keep evaluation defensible, teams often rely on established quality and safety frameworks. For general chemical safety and handling expectations, SDS guidance is commonly aligned with recognized regulatory systems, such as the Globally Harmonized System of Classification and Labelling of Chemicals (GHS). For quality systems and documentation rigor, many organizations follow principles consistent with pharmaceutical quality guidance and related industry best practices. For general principles, consult:
If your use-case is regulated (for example, pharmaceutical or clinical manufacturing), you should apply the relevant national regulations and internal quality standards in addition to any supplier documents for Pmoc Abrava. In practice, “objective background” means your decisions should be auditable: you can show why you selected the supplier, what evidence you relied on, and what checks you performed to ensure ongoing suitability.
Beyond regulatory guidance, many professional teams also rely on internal standard operating procedures (SOPs) for receiving, handling, sampling, and storage. Where feasible, teams incorporate trending—comparing incoming CoA values over time and tracking whether any measured impurity levels drift upward even if still within spec. Trending is particularly useful for intermediates where subtle shifts can precede performance changes.
Pmoc Abrava is referenced as a specific chemical input in some technical contexts. The exact identity and role can vary depending on how it is named in a given dataset or supplier documentation. For accurate determination, confirm the molecular identity, CAS number (if applicable), and the supplier’s specification sheet for the exact product offering. In professional settings, relying on internal shorthand names without verifying the full identity is a common root cause for mis-procurement or misqualification.
Compare documentation and qualification readiness: request lot-specific CoA, review impurity profiles and test methods, verify storage guidance, and confirm traceability and change control practices. If possible, conduct a small-scale suitability test under your process conditions. In addition, consider whether suppliers provide CoA history across multiple lots or at least enough data to understand typical variability. Variability itself is a key parameter in process robustness.
Price matters, but it is usually secondary to technical fit and risk. If a batch fails acceptance or causes purification changes, the total cost can exceed savings from a lower unit price. A cost-of-quality approach often produces more reliable outcomes. In mature organizations, this is formalized as expected cost per successful batch or cost per unit of usable intermediate/purified output—not just the purchase price.
Follow the SDS and supplier-specific guidance for storage, moisture protection, and safe handling. In general, disciplined weighing, contamination control, and correct solvent compatibility checks are critical to reduce variability for intermediate-like inputs such as Pmoc Abrava. Additionally, implement training and checklists so that routine operations do not gradually drift away from qualified practices.
Incoming checks typically include container/label verification, lot identity confirmation, and any identity or purity screening required by your internal QA plan. If your process is sensitive, add a technical suitability verification to confirm batch performance. The goal is to prevent defective materials from entering production runs where they would consume capacity and potentially compromise schedule-critical deliverables.
Expectations depend on your end-use and jurisdiction. If used in regulated production, quality system controls (documentation, traceability, change control, and audit readiness) are commonly required. Use your organization’s compliance framework and the supplier’s documentation to confirm readiness for your specific context. Even if not explicitly regulated, many organizations adopt similar practices to improve reliability and reduce operational risk.
Interchangeability should not be assumed. Differences in impurity profiles, physical form, or manufacturing details can affect performance. Qualification should be performed for new lots and new suppliers based on your acceptance criteria. Some organizations implement “light requalification” for certain changes (for example, minor CoA shifts within typical variability), while reserving full requalification for major changes. The key is defining triggers objectively.
For professionals working with Pmoc Abrava, the very effective strategy is to treat it as a process parameter, not merely a commodity. Start with a clearly defined specification need, require lot-specific documentation, qualify performance through small-scale testing, and enforce stable handling and traceability. This approach reduces uncertainty, protects timelines, and supports defensible quality decisions—whether your work is in development, pilot production, or a broader manufacturing program.
To further tighten the workflow, many teams implement a “feedback loop” where process outcomes inform future qualification depth and acceptance criteria. For example, if incoming impurity trends correlate with purification strain, you can adjust what you monitor or increase internal verification frequency. This turns supplier qualification into an evolving program rather than a one-time gate.
If you share the actual context you’re working in (e.g., the chemical’s full name as shown on your SDS/spec sheet, intended reaction step, and your target purity/impurity constraints), I can tailor the qualification checklist and decision criteria more precisely for Pmoc Abrava.
Striking the Perfect Balance: Navigating Premiums and Out-of-Pocket Expenses in Senior Insurance Plans

Explore the Tranquil Bliss of Idyllic Rural Retreats

How to Make Lasting Memories at Disneyland Attractions

4rkojvsx Technology: Industry Insights and Buying Guide

Affordable Phones and Plans for Seniors

Affordable Full Mouth Dental Implants Near You

Unlock the Top Kept Secrets to Finding Your Ideal Dentist for Flawless Dental Implant Results!

Discovering Springdale Estates

Unveiling RS Sul Telecom Services

Pet Insurance Essentials: Coverage, Costs, and Resources Explained
In recent years, pet ownership has seen a significant surge. Pets, whether they're dogs, cats, birds, or other animals, have increasingly become integral parts of our families. With this growing bond, there is a rising awareness of the importance of pet health and well-being. Just as health insurance is crucial for humans, pet insurance is becoming an essential consideration for pet owners. This article aims to provide a comprehensive understanding of pet insurance, covering its importance, types of coverage, associated costs, and valuable resources to help you make an informed decision.
Colbate Explained: Uses, Sourcing, and Top Practices
This guide explains Colbate from a practical industry viewpoint, covering what it is, how suppliers typically handle it, and what buyers should verify before purchase. It also summarizes objective background on related terminology, common application considerations, and due-diligence steps. Readers will find a comparison table, requirements, and targeted FAQs to support confident sourcing decisions.
Strategies for Financial Security and Legacy Planning
Life insurance isn't just about what happens after you pass—it's a strategic tool in a comprehensive plan for your financial security and estate planning.
Navigating Insurance Costs for Seniors: Essential Tips and Advice
As we age, healthcare costs and the complexity of managing them can become increasingly burdensome. For many seniors, navigating insurance costs is a crucial aspect of ensuring financial stability and maintaining health. This guide aims to provide you with essential tips and advice on how to manage and potentially reduce these costs effectively.
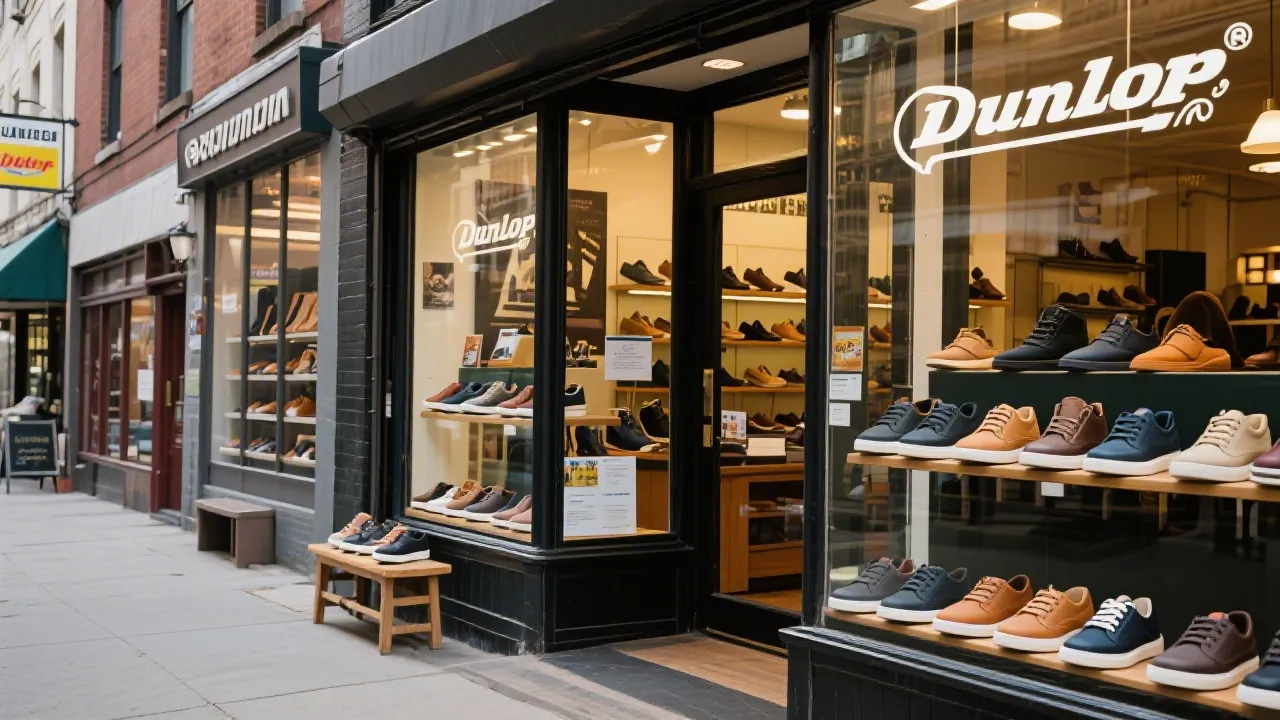
Discovering Dunlop Brooklin Footwear
This article delves into Dunlop Brooklin, a prominent name in the footwear industry known for its exceptional quality and innovation. Dunlop is recognized for its wide-ranging collection of shoes designed to meet diverse customer needs, from casual wear to professional attire. This guide provides an in-depth analysis of Dunlop Brooklin and its offerings, revealing why it stands out in the market.
Your Guide to Understanding Car Insurance Coverage
Navigating car insurance can be puzzling. Fortunately, this guide illuminates coverage types and choosing the right provider.
